
endast för forskning
XL147 analogue PI3K Hämmare
Kat.nr.: S1118
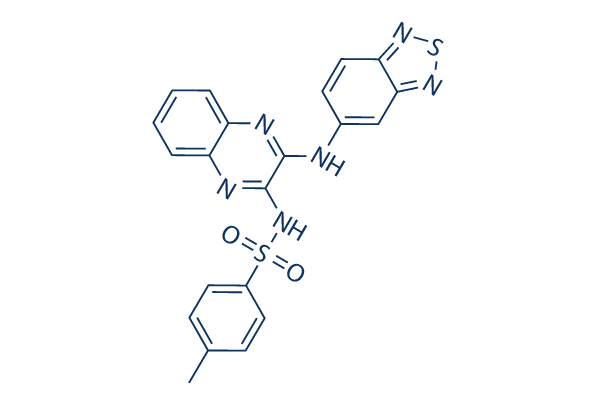
Kemisk struktur
Molekylvikt: 448.52
Kvalitetskontroll (Quality Control)
| Relaterade mål | Akt mTOR GSK-3 ATM/ATR DNA-PK AMPK PDPK1 PTEN PP2A PDK |
|---|---|
| Övrigt PI3K Inhibitorer | GDC-0077 (Inavolisib) SAR405 Quercetin (Sophoretin) LY294002 Tersolisib (STX-478) Buparlisib (BKM120) 740 Y-P (PDGFR 740Y-P) GO-203 TFA Eganelisib (IPI-549) Paxalisib (GDC-0084) |
Kemisk information, lagring & stabilitet (Chemical Information, Storage & Stability)
| Molekylvikt | 448.52 | Formel | C21H16N6O2S2 |
Lagring (från mottagningsdatumet) | |
|---|---|---|---|---|---|
| CAS-nr | 956958-53-5 | Ladda ner SDF | Lagring av stamlösningar |
|
|
| Synonymer | SAR245408 | Smiles | CC1=CC=C(C=C1)S(=O)(=O)NC2=NC3=CC=CC=C3N=C2NC4=CC5=NSN=C5C=C4 | ||
Löslighet (Solubility)
|
In vitro |
DMSO
: 3 mg/mL
(6.68 mM)
Water : Insoluble Ethanol : Insoluble |
Molaritetsräknare
|
In vivo |
|||||
In vivo-formuleringsräknare (Klar lösning)
Steg 1: Ange information nedan (Rekommenderas: Ett extra djur för att ta hänsyn till förluster under experimentet)
Steg 2: Ange in vivo-formuleringen (Detta är endast räknaren, inte formuleringen. Vänligen kontakta oss först om det inte finns någon in vivo-formulering i löslighetsavsnittet.)
Beräkningsresultat:
Arbetskoncentration: mg/ml;
Metod för att bereda DMSO-stamlösning: mg substans förlöst i μL DMSO ( Stamlösningskoncentration mg/mL, Vänligen kontakta oss först om koncentrationen överskrider DMSO-lösligheten för denna batch av substansen. )
Metod för att bereda in vivo-formulering: Ta μL DMSO stamlösning, tillsätt sedanμL PEG300, blanda och klarna, tillsätt sedanμL Tween 80, blanda och klarna, tillsätt sedan μL ddH2O, blanda och klarna.
Metod för att bereda in vivo-formulering: Ta μL DMSO stamlösning, tillsätt sedan μL Majsolja, blanda och klarna.
Obs: 1. Se till att vätskan är klar innan du tillsätter nästa lösningsmedel.
2. Se till att tillsätta lösningsmedlet/lösningsmedlen i ordning. Du måste se till att lösningen som erhålls i föregående tillsats är en klar lösning innan du fortsätter att tillsätta nästa lösningsmedel. Fysiska metoder som vortex, ultraljud eller varmt vattenbad kan användas för att underlätta upplösningen.
Verkningsmekanism (Mechanism of Action)
| Targets/IC50/Ki |
PI3Kγ
(Cell-free assay) 23 nM
PI3Kδ
(Cell-free assay) 36 nM
PI3Kα
(Cell-free assay) 39 nM
PI3Kβ
(Cell-free assay) 383 nM
|
|---|---|
| In vitro |
XL147 analogue inhibits class I PI3K isoforms in an ATP-competitive manner. In a panel of HER2-overexpressing human breast cancer cell lines, treatment with this compound abrogates AKT and S6 phosphorylation but also induces the expression and phosphorylation of HER3 and other RTKs. In HER2+ cells, phosphorylation of HER3 is maintained by the HER2 tyrosine kinase, leading to partial recovery of phosphorylated AKT (pAKT) and thereby limiting the antitumor action of this chemical. In addition, knockdown of HER3 or treatment with the anti-HER2 agents trastuzumab or lapatinib sensitizes HER2+ breast cancer cells to this agent in vitro and in vivo. Treatment with this inhibitor inhibits the monolayer growth of all tested cell lines, including BT474, HCC1937 et al. in a dose-dependent manner. The main effect of this compound is inhibition of cell proliferation. It induces cell death at the concentration of 20 μM. Treatment with this chemical leads to dose-dependent inhibition of PI3K. Consistent with the inhibition of cell proliferation, it induces a reduction in cyclin D1 and pRB and an increase in levels of the CDK inhibitor p27KIPI but no detectable change in levels off total or cleaved poly (ADP-ribose) polymerase (PARP). Treatment with this agent leads to a dose-dependent reduction in pAKTS473/T308 and pS6S240/244. Surprisingly, it also triggers up-regulation of total HER3 and/or pHER3Y1289 levels. In HER2-overexpressing cells, inhibition of PI3K is followed by up-regulation of expression and phosphorylation of multiple receptor tyrosine kinases, including HER3. Knockdown of FoxO1 and FoxO3a transcription factors prevents the induction of HER3, InsR, IGF1R, and FGFR2 mRNAs upon inhibition of PI3K. In HER2+ cells, knockdown of HER3 with siRNA or cotreatment with the HER2 inhibitors trastuzumab or lapatinib enhances XL147-induced cell death and inhibition of pAKT and pS6.
|
| In vivo |
Athymic mice with BT474 xenografts are randomly treated with XL147 analogue, lapatinib, trastuzumab, or this compound plus each HER2 antagonist. Each monotherapy significantly inhibtis tumor growth with trastuzumab being the only agent that induced a complete tumor regression in one of eight mice. Both combinations are superior to the respective drugs given alone. Notably, the combination of trastuzumab and this chemical, but not lapatinib and XL147, induces a complete tumor response in three of eight mice. There is no marked drug-related toxicity in any of the treatment arms. The combination of this compound plus trastuzumab prevents pHER3 more potently than any of the other treatments. In good agreement with differences in tumor growth among treatment arms, nuclear pAKT is lower in tumors treated with XL147 plus lapatinib or this chemical plus trastuzumab compared with tumors treated with single agents. Of all three single drugs, this compound is the only one shown statistically to repress nuclear pAKT levels. There are no detectable changes in cytoplasmic pAKT levels. Combined inhibition of HER2 and PI3K in HER2-dependent xenografts is required to maximally inhibit signaling output of the PI3K/AKT pathway.
|
Referenser |
|
Klinisk prövningsinformation (Clinical Trial Information)
(data från https://clinicaltrials.gov, uppdaterad den 2024-05-22)
| NCT-nummer | Rekrytering | Tillstånd | Sponsor/Samarbetspartners | Startdatum | Faser |
|---|---|---|---|---|---|
| NCT01943838 | Completed | Neoplasm Malignant |
Sanofi |
October 2013 | Phase 1 |
| NCT01436565 | Completed | Solid Tumor Cancers |
Sanofi|Merrimack Pharmaceuticals |
November 2011 | Phase 1 |
| NCT01392924 | Completed | Neoplasm Malignant |
Sanofi |
August 2011 | Phase 1 |
| NCT01357330 | Completed | Solid Tumors |
Sanofi |
May 2011 | Phase 1 |
| NCT01240460 | Completed | Glioblastoma|Astrocytoma Grade IV |
Sanofi |
January 2011 | Phase 1 |

Produkterna är endast avsedda för forskning. Ej för användning på människor. Vi säljer inte till patienter.
©Copyright 2013 Selleck Chemicals. Med ensamrätt.